rDock is an open source docking program, trying to solve the same kind of scientific questions as Autodock Vina, which I have covered in a couple of
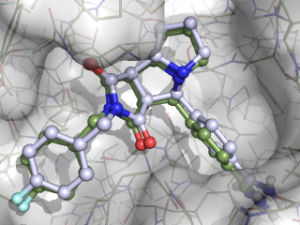
rDock is an open source docking program, trying to solve the same kind of scientific questions as Autodock Vina, which I have covered in a couple of
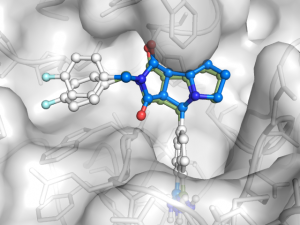
In the two previous blog posts Ligand docking with Smina and Never use re-docking for …, it was demonstrated how easy it is to dock a small ligand
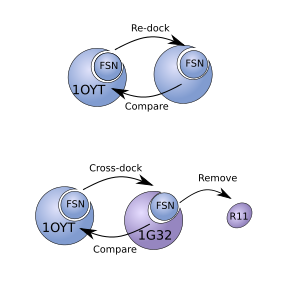
Re-docking of ligands found in PDB files are often used as a fast evaluation of a docking program before working with designed or other ligands. However this
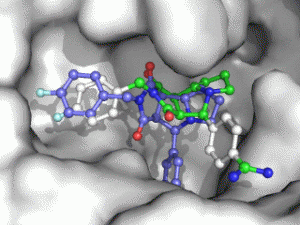
Moleculer Docking is a powerful technique for studying potential ligand-receptor interactions. It can be done with free tools. In this blog post I showcase the docking program
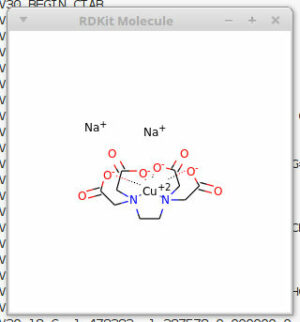
When I have been working with chemical databases and import of molecules I have encountered numerous problems with the way chemical structures are drawn. Most often the
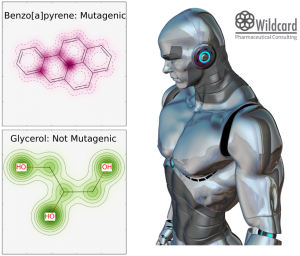
Toxic compounds are most often something that we try to avoid when designing novel pharmaceutical compounds, so it could be nice to get a prediction if a
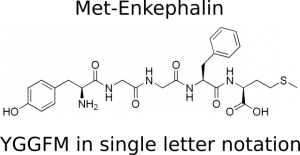
The chemoinformatics package Rdkit has is strength with handling small organic molecules. These molecules are characterized by a large diversity in chemical structures. A description of the
Feature selection is a powerful way of reducing the complexity of a machine learning or statistical model. But feature selection must be done in the right way,
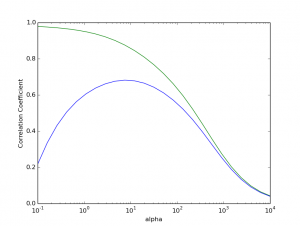
Last time a simple multiple linear regression (MLR) model was seriously overfitted to molecular solubility data. This time the concept of regularization will be tested. Recall that
Last blog entry the conversion between molecule and fingerprint was briefly touched upon. Now the fingerprints will be used as the basis for a simple attempt to